Water: Energy, Orbitals, and Charges
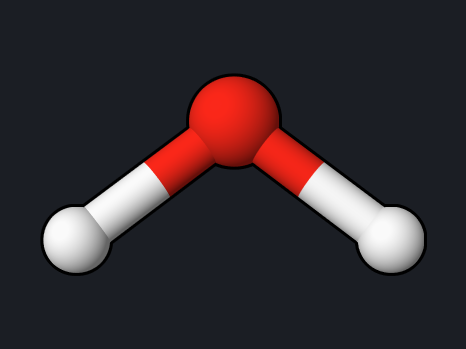
A single-point DFT calculation is the simplest OpenQP run. Here it gives the total energy of water, the energy of every molecular orbital, pictures of the frontier orbitals, the dipole moment, and Mulliken atomic charges.
Copy and run
dft/pbe0/6-31g* properties(scf_prop="el_mom,mulliken")
geom="""
O      0.000000     0.000000     0.122745
H      0.000000     0.759541    -0.471922
H      0.000000    -0.759541    -0.471922
"""
openqp water-scf.oqp
The first line chooses DFT with the PBE0 functional and the 6-31G* basis set.
No task keyword means a single-point energy. properties(scf_prop=...)
requests the dipole moment (el_mom) and Mulliken charges (mulliken); without
it, OpenQP prints only the energy and orbitals. The coordinates are in ångström.
This run takes about 2 seconds.
What you get
| Quantity | Reference value |
|---|---|
| Total energy | −76.3256994 hartree |
| SCF iterations | 8 |
| Point group detected | C2v |
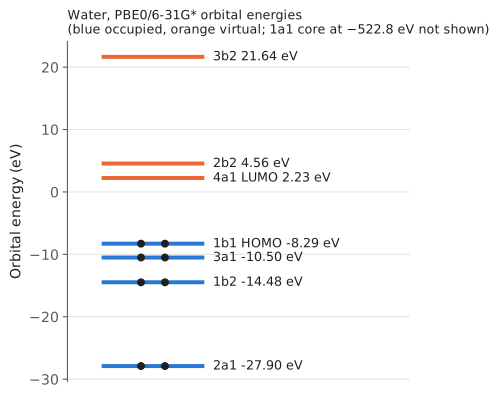
| HOMO energy (1b1) | −0.30453 hartree (−8.29 eV) |
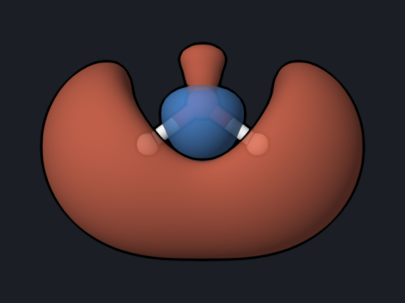
| LUMO energy (4a1) | 0.08193 hartree (2.23 eV) |
| HOMO–LUMO gap | 10.52 eV |
| Dipole moment | 2.125 debye |
| Mulliken charges | O −0.803, H +0.402 |
The SCF section of water-scf.log ends with the converged energy. OpenQP
prints RHF for any closed-shell reference, including Kohn–Sham DFT:
Final RHF energy is -76.3256993935 after 8 iterations
OpenQP labels every orbital with its symmetry in the detected point group:
   point group:      c2v
   alpha MOs:
       1      -19.21076550  a1
       2       -1.02527488  a1
       3       -0.53207181  b2
       4       -0.38604085  a1
       5       -0.30452890  b1
       6        0.08193402  a1
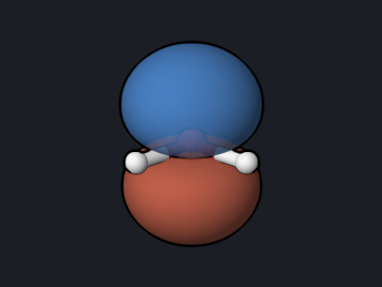
Orbitals 1–5 are occupied (10 electrons). Orbital 1 is the oxygen 1s core orbital, orbitals 2–4 are the O–H bonding and in-plane lone-pair orbitals, and orbital 5, the HOMO, is the out-of-plane oxygen lone pair. The dipole moment and charges follow:
    electric dipole (Debye):
                                                   X              Y              Z             Norm
                                         D_o    -0.00000000    -0.00000000    -2.12548100     2.12548100

       #    Name        Charge
------------------------------
       1    O        -0.803453
       2    H         0.401727
       3    H         0.401727
The dipole lies along the C2 axis (z), as symmetry requires. The value is larger than the experimental gas-phase dipole of water (1.85 D), which is typical of a small basis set without diffuse functions. Mulliken charges depend strongly on the basis set and are useful for trends, not as absolute values.
The SCF iterations
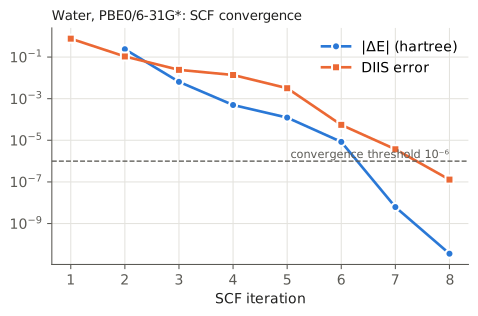
The energy change and the DIIS error fall by about an order of magnitude per iteration once DIIS takes over at iteration 2; the run stops when the error is below the threshold of 10⁻⁶.
All valence orbitals
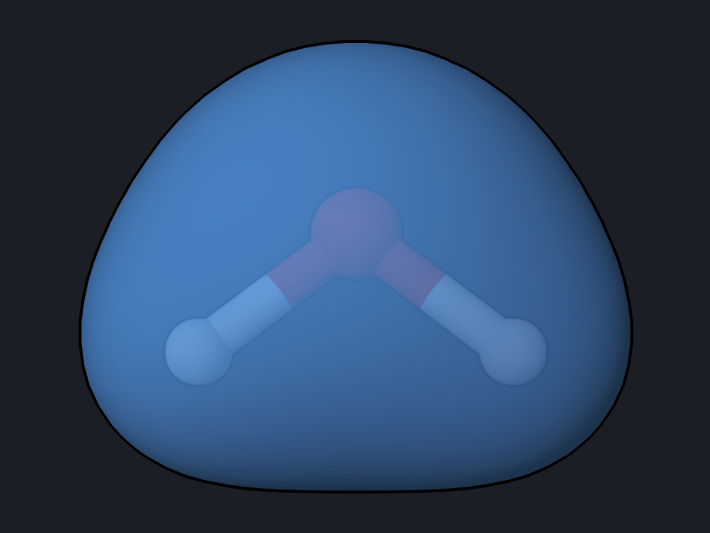
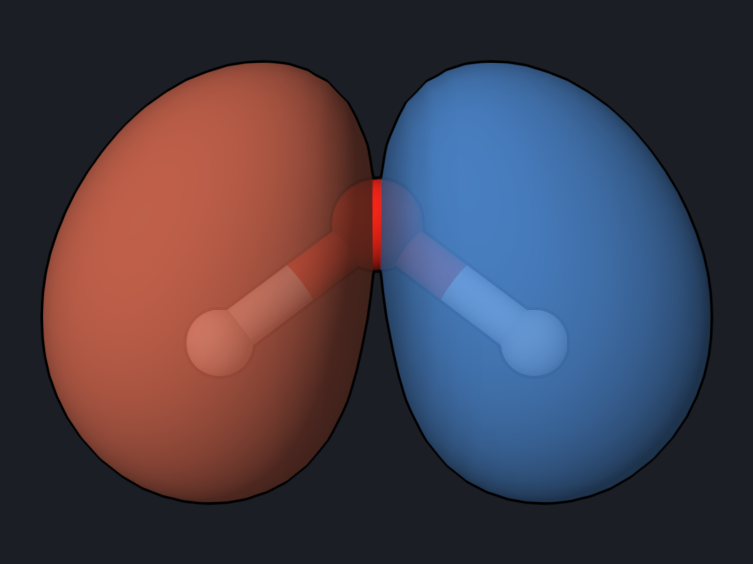
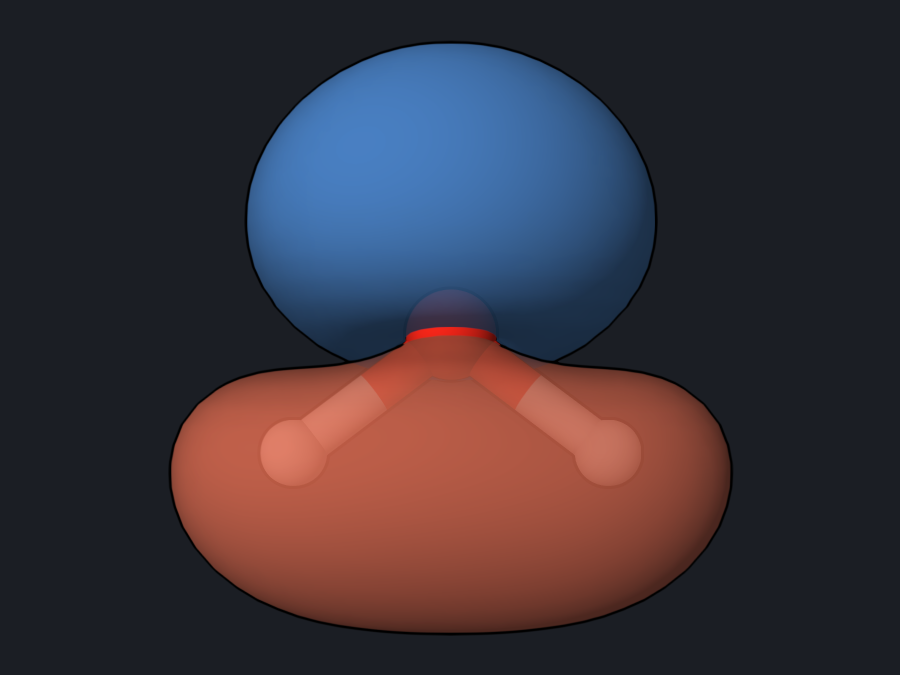
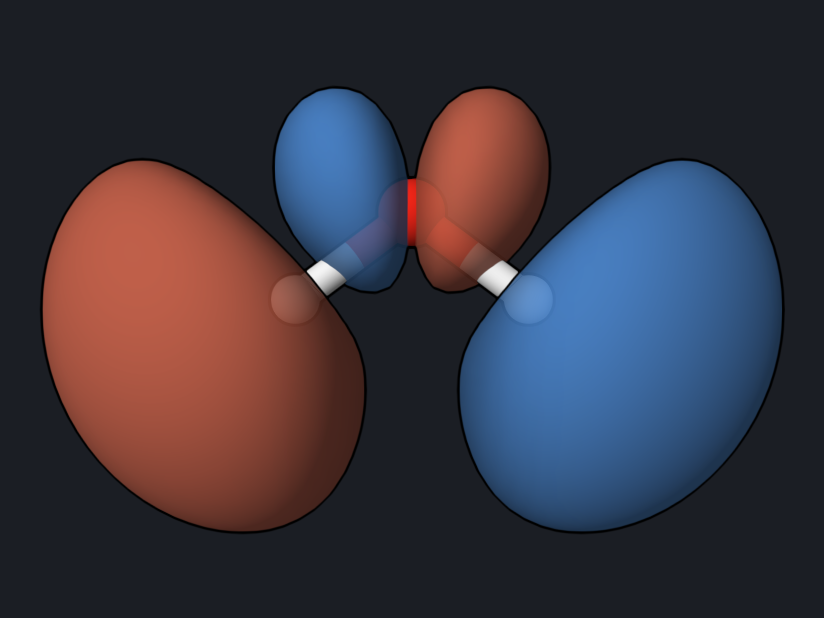
Look at the orbitals
The orbitals are in water-scf_scf_rhf_pbe0_6-31gs.molden. Open this file in
OQP Studio Analysis or in
OpenqpView and select
orbital 5 (HOMO) or 6 (LUMO). The pictures above use an isovalue of
0.05 a.u.; red and blue mark the two signs of the orbital.
Atomic charges on a larger molecule
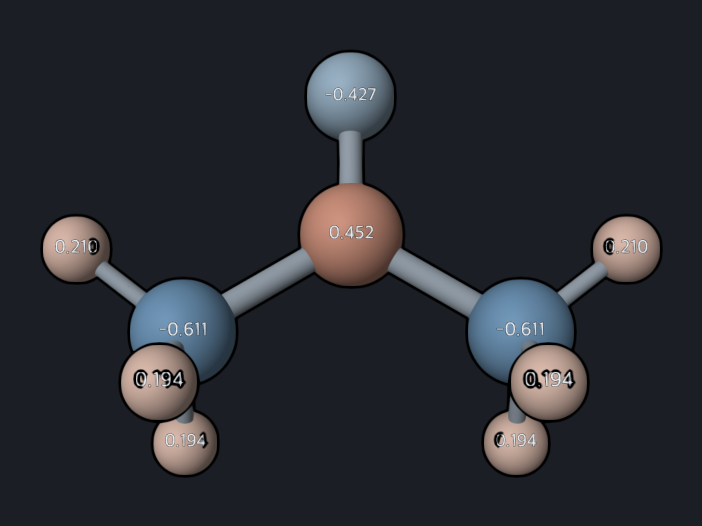
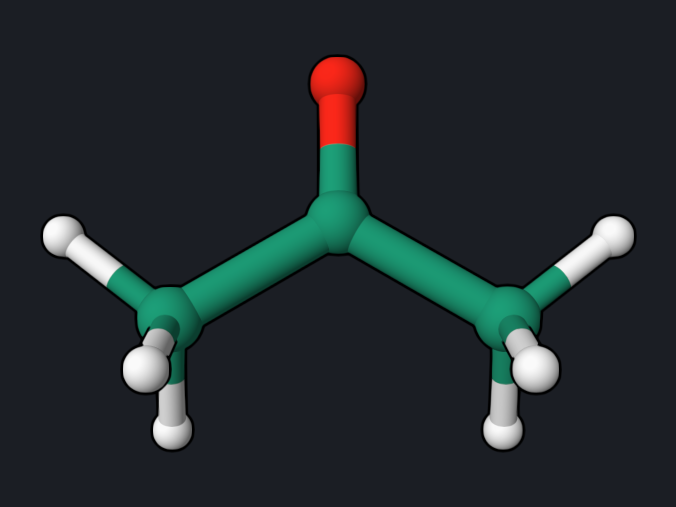
The same properties(...) call works for any molecule. For acetone at its
PBE0/6-31G* geometry (acetone-charges.oqp), the
dipole moment is 2.856 D and the Mulliken charges are O −0.427, carbonyl C
+0.452, methyl C −0.611, and H +0.194 to +0.210. OQP Studio can color the
atoms by these charges and print each value on its atom:
Try next
- Change
6-31g*tocc-pvtz: the dipole moment drops to 1.93 D, closer to experiment. - Replace
dft/pbe0withhffor a Hartree–Fock calculation; the HOMO–LUMO gap grows from 10.5 eV to 19.2 eV. - Add
lowdintoscf_propfor Löwdin charges, for examplescf_prop="el_mom,mulliken,lowdin".
See also: HF and DFT,
properties.
Scientific references
HF and density functional theory; Exchange-correlation functionals.