LiH Dissociation: CASSCF, CASPT2, and FCI
CASPT2 adds second-order perturbation theory on top of a CASSCF reference: CASSCF handles the near-degeneracy of a breaking bond, and CASPT2 adds the remaining dynamic correlation. LiH is small enough for full configuration interaction (FCI), the exact answer in a given basis set, so every method on this page can be checked against it.
Copy and run
caspt2/6-31g cas(active_electrons=2,active_orbitals=2,frozen_core=1,max_det=200000)
geom="""
Li 0.000000 0.000000 0.000000
H 0.000000 0.000000 1.600000
"""
openqp lih-caspt2.oqp
caspt2/6-31g: single-state CASPT2 on a CASSCF reference.cas(active_electrons=2,active_orbitals=2,frozen_core=1,...): two electrons in the σ bonding and σ* antibonding orbitals; the Li 1s pair is frozen in the perturbation step.max_det=200000: OpenQP's CASPT2 is uncontracted, so its first-order space is a set of determinants.max_detcaps its size; the default of 5000 is also enough for this system (the run gives the same energy without it), and OpenQP stops with a message naming the required size when a system needs more. Uncontracted CASPT2 is meant for small molecules and active spaces; for ethylene in 6-31G* the space would already hold about 4 × 10¹² determinants.frozen_coreincas(...)is the number of doubly occupied orbitals outside the active space (here the Li 1s pair). OpenQP checks it againstactive_electrons: for ethylene with CAS(2,2),frozen_core=2is rejected withfrozen_core is inconsistent with active_electrons, andfrozen_core=7(all seven doubly occupied orbitals) is accepted.
The same geometry with the other methods (each run takes 1–2 seconds):
| Method | First line | Download |
|---|---|---|
| RHF | rhf/6-31g |
lih-rhf.oqp |
| UHF | uhf/6-31g scf(stability=true) |
lih-uhf.oqp |
| CASSCF(2,2) | casscf/6-31g cas(active_electrons=2,active_orbitals=2,frozen_core=1) |
lih-casscf.oqp |
| CASPT2(2,2) | as above | lih-caspt2.oqp |
| CCSD(T) | ccsd_t/6-31g |
lih-ccsdt.oqp |
| FCI | fci/6-31g fci(max_det=200000) |
lih-fci.oqp |
For the curve, change the H coordinate (the Li–H distance, in Å) from 1.0 to 5.0 Å.
What you get
The CASPT2 block of the log shows the reference, the size of the first-order space, and the second-order energy:
   PyOQP active space:                 CAS(2,2)
   PyOQP PT2 frozen-core orbitals:     1
   PyOQP external (first-order) space:  96
   PyOQP E(0) reference  (CASSCF):          -7.9959166652
   PyOQP E(2) correlation:                  -0.0017465707
   PyOQP E(CASPT2  ) total:             -7.9976632359
The FCI run reports PyOQP determinant count: 3025: all 4 electrons in all
11 orbitals of 6-31G.
| Method | 1.6 Å (hartree) | 5.0 Å (hartree) | Error vs FCI at 1.6 Å |
|---|---|---|---|
| RHF | −7.97932 | −7.82238 | +19.0 mEh |
| UHF | −7.97932 | −7.92966 | +19.0 mEh |
| CASSCF(2,2) | −7.99592 | −7.92992 | +2.4 mEh |
| CASPT2(2,2) | −7.99766 | −7.92993 | +0.7 mEh |
| CCSD(T) | −7.99836 | −7.93046 | 0.0 mEh |
| FCI | −7.99836 | −7.93025 | — |
Reading the result
- RHF dissociates to the wrong limit (ionic Li⁺H⁻ character), as for
H₂. UHF with
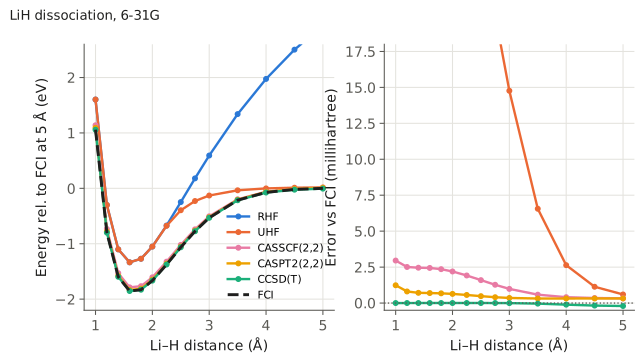
scf(stability=true)breaks spin symmetry beyond about 2.25 Å and then reaches the right limit, but is 19 mEh too high near equilibrium. - CASSCF(2,2) has the right shape everywhere: the error is 2.4 mEh at 1.6 Å and falls to 0.3 mEh at 5 Å.
- CASPT2(2,2) reduces the error at equilibrium to 0.7 mEh. Part of what remains is the correlation of the frozen Li 1s pair, which FCI includes.
- CCSD(T) is essentially exact near equilibrium for this four-electron system. At 5 Å it is 0.2 mEh below FCI: CCSD(T) is not variational, and its perturbative triples become less reliable as the bond breaks.
- The FCI dissociation energy in this basis is 0.06811 hartree (1.85 eV), from the energies at 1.6 and 5.0 Å.
Optimize the bond length with analytic gradients
CASSCF, CASPT2, and strongly contracted NEVPT2 (SC-NEVPT2) all have analytic
nuclear gradients in OpenQP, so opt works directly. For SC-NEVPT2, the
zeroth-order Hamiltonian and contraction are selected in pt2(...):
caspt2/6-31g opt cas(active_electrons=2,active_orbitals=2,frozen_core=1) pt2(h0=dyall,contraction=strong,gradient=analytic)
geom="""
Li     0.000000     0.000000     0.000000
H      0.000000     0.000000     1.600000
"""
Download lih-nevpt2-opt.oqp ·
lih-caspt2-opt.oqp (first line
caspt2/6-31g opt cas(...) pt2(gradient=analytic)) ·
lih-casscf-opt.oqp
pt2(h0=dyall,contraction=strong) turns the CASPT2 route into SC-NEVPT2;
gradient=analytic requests the analytic gradient and stops with an error
rather than silently falling back to finite differences if a setting is not
covered. The log confirms the route (Analytic SC-NEVPT2 Nuclear Gradient),
and each optimization takes 2–7 seconds.
| Method | r(Li–H) |
|---|---|
| CASSCF(2,2), analytic gradient | 1.6736 Å |
| CASPT2(2,2), analytic gradient | 1.6721 Å |
| SC-NEVPT2(2,2), analytic gradient | 1.6727 Å |
| CCSD(T), fitted to a 0.025 Å scan | 1.6717 Å |
| FCI, fitted to a 0.025 Å scan | 1.6717 Å |
All three multireference bond lengths lie within 0.002 Å of FCI in this basis set. (The experimental bond length, 1.596 Å, needs a much larger basis set.)
Try next
- Replace
caspt2withxms-caspt2and addci(nroot=2,target_spin=singlet) pt2(target_roots="0,1",nroot=2)to treat the first excited singlet as well. At 1.6 Å this gives −7.99231 and −7.87520 hartree for the two states, an excitation energy of 3.19 eV. - Replace
caspt2withmrmp2: for this single-state CAS(2,2) case it gives the same energy as CASPT2, −7.9976632 hartree.
See also: CASPT2 Nuclear Gradient, Wavefunction methods, H₂ dissociation.