Benzene: NMR Chemical Shifts
An NMR calculation gives the absolute magnetic shielding σ of every nucleus. Spectra report chemical shifts δ relative to a reference compound, so two calculations at the same level are needed: the molecule of interest and the reference, tetramethylsilane (TMS). Then
δ = σ(TMS) − σ(molecule).
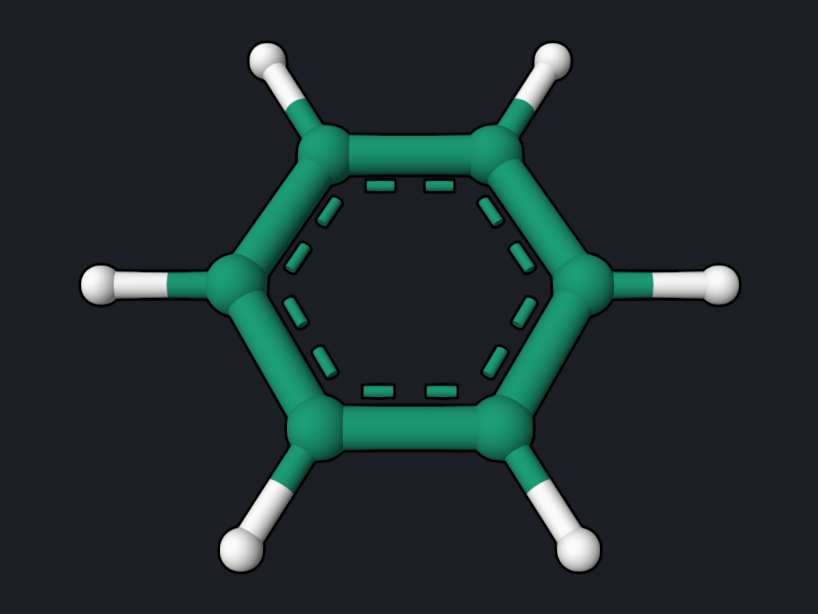
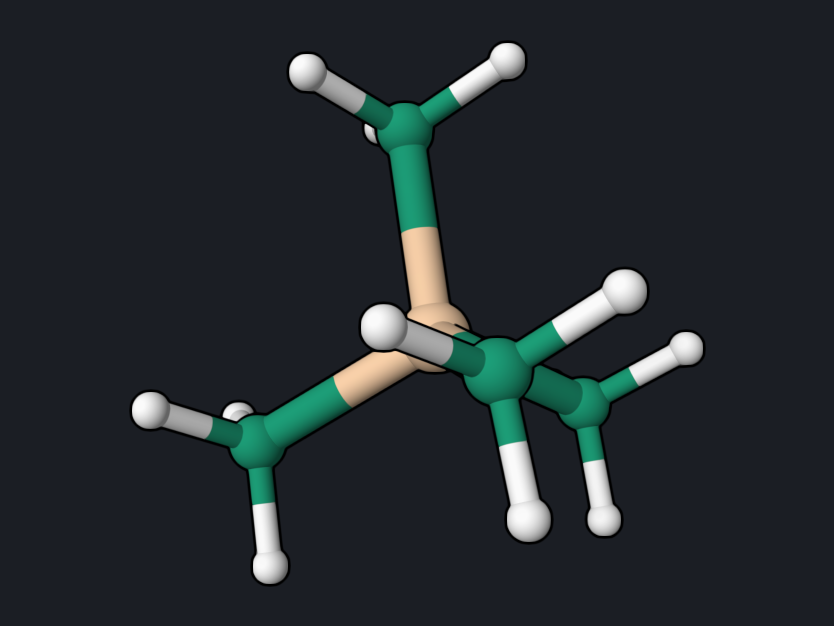
Copy and run
dft/pbe0/6-31g* nmr
geom="""
C      1.392883     0.000000     0.000000
C      0.696465     1.206259     0.000000
C     -0.696465     1.206259     0.000000
C     -1.392883     0.000000     0.000000
C     -0.696465    -1.206259     0.000000
C      0.696465    -1.206259     0.000000
H      2.479666     0.000000     0.000000
H      1.239913     2.147463     0.000000
H     -1.239913     2.147463     0.000000
H     -2.479666     0.000000     0.000000
H     -1.239913    -2.147463     0.000000
H      1.239913    -2.147463     0.000000
"""
Download benzene-nmr.oqp · Download tms-nmr.oqp (TMS, 17 atoms, same first line)
openqp benzene-nmr.oqp
openqp tms-nmr.oqp
nmr computes GIAO (gauge-including atomic orbital) shieldings, which do not
depend on the choice of coordinate origin. Both geometries were optimized at
PBE0/6-31G*. Benzene takes about 25 seconds and TMS about 40 seconds.
What you get
The end of each log has the isotropic shielding of every atom, in ppm. The
column to use is total_coupled, which includes the response of the orbitals to
the magnetic field (coupled-perturbed Kohn–Sham). For benzene:
    Isotropic shielding (GIAO, ppm)   [exact-exchange c_x =  0.2500]
       Atom    Z   sigma_dia   para_uncoupled   para_coupled   total_uncoupled   total_coupled
         1   6.0      256.824629     -143.610570     -184.723650      113.214059       72.100980
         2   6.0      256.835108     -143.625026     -184.741975      113.210082       72.093133
       ...
         7   1.0       29.549782       -3.924546       -4.810611       25.625236       24.739171
         8   1.0       29.545189       -3.921758       -4.807423       25.623431       24.737766
OQP Studio can color the atoms by these values and print them on the structure:
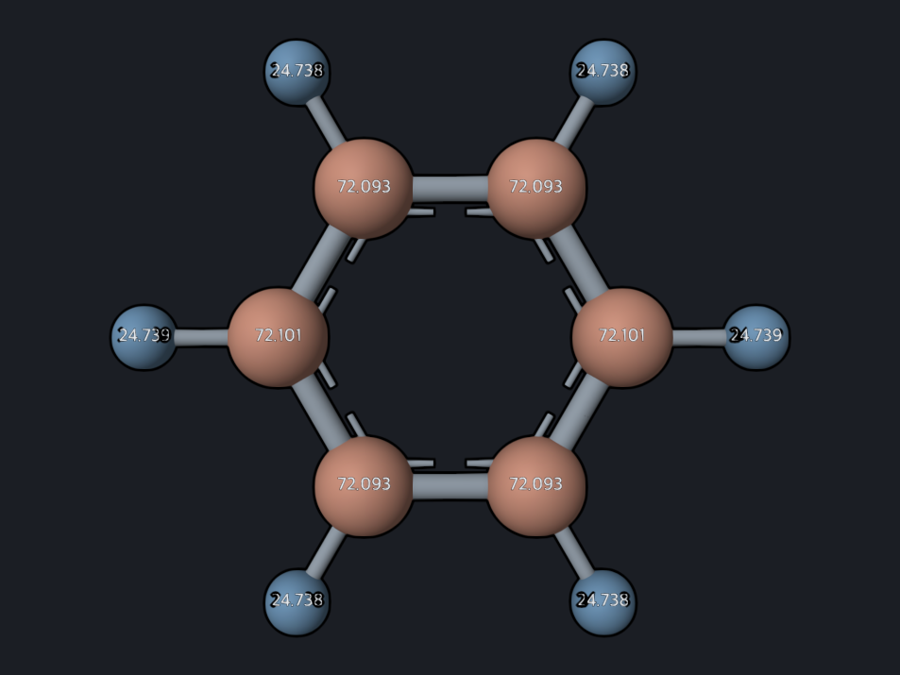
Equivalent atoms agree to 0.01 ppm; the small spread comes from the DFT integration grid. Averaging over equivalent atoms:
| Nucleus | σ(benzene) | σ(TMS) | δ = σ(TMS) − σ | Experiment (CDCl₃) |
|---|---|---|---|---|
| ¹H | 24.738 ppm | 32.109 ppm | 7.37 ppm | 7.34 ppm |
| ¹³C | 72.10 ppm | 194.49 ppm | 122.4 ppm | 128.4 ppm |
TMS has four equivalent carbons and twelve equivalent hydrogens; its geometry here is not exactly Td, and the individual values spread by 0.2 ppm (C) and 0.02 ppm (H) around the averages above.
Reading the result
The ¹H shift is close to experiment. The ¹³C shift is 6 ppm too low; ¹³C
shieldings depend more strongly on the functional and the basis set than ¹H
shieldings, so test a larger basis set such as cc-pvtz before relying on
absolute ¹³C values. The experimental values are for solution, while the
calculation is for an isolated molecule.
Cross-check
To test the GIAO implementation, the benzene calculation was repeated at the
Hartree–Fock level (hf/6-31g* nmr) in OpenQP and in PySCF 2.13 at the same
geometry. Both give σ(C) = 73.519 ppm and σ(H) = 25.083 ppm.
Try next
- Compute the shifts of a substituted benzene, such as toluene, at the same level; the TMS reference values above can be reused.
- Repeat the benzene run at the Hartree–Fock level,
hf/6-31g* nmr, and compare with the values in the cross-check above.
See also: NMR, IR, and Raman,
properties.